
Recall update: FDA now urging clinicians to return vascular device linked to 3 deaths
The U.S. Food and Drug Administration (FDA) has shared updated recommendations about recalled thoracic stent grafts. Customers were previously told they could still use the devices as long as they were aware of the risk and knew how to respond if an issue occurred. Now, however, the FDA is emphasizing that these devices should be returned to the manufacturer and not be used to treat any other patients.
Background details about this critical issue
Back in April, the FDA warned the public about an ongoing safety issue with certain Relay Pro Thoracic Stent Grafts from Terumo Aortic.
These grafts, used by interventional cardiologists and vascular surgeons to repair damaged or weakened sections of a patient’s thoracic aorta, were malfunctioning and failing to unclasp from the delivery system. The issue was linked to an increased risk of procedure delays and stent graft displacement; surgical repair procedures can even be necessary in certain instances.
The issue was linked to a total of three patient deaths, but customers were not told to return the devices to the manufacturer.
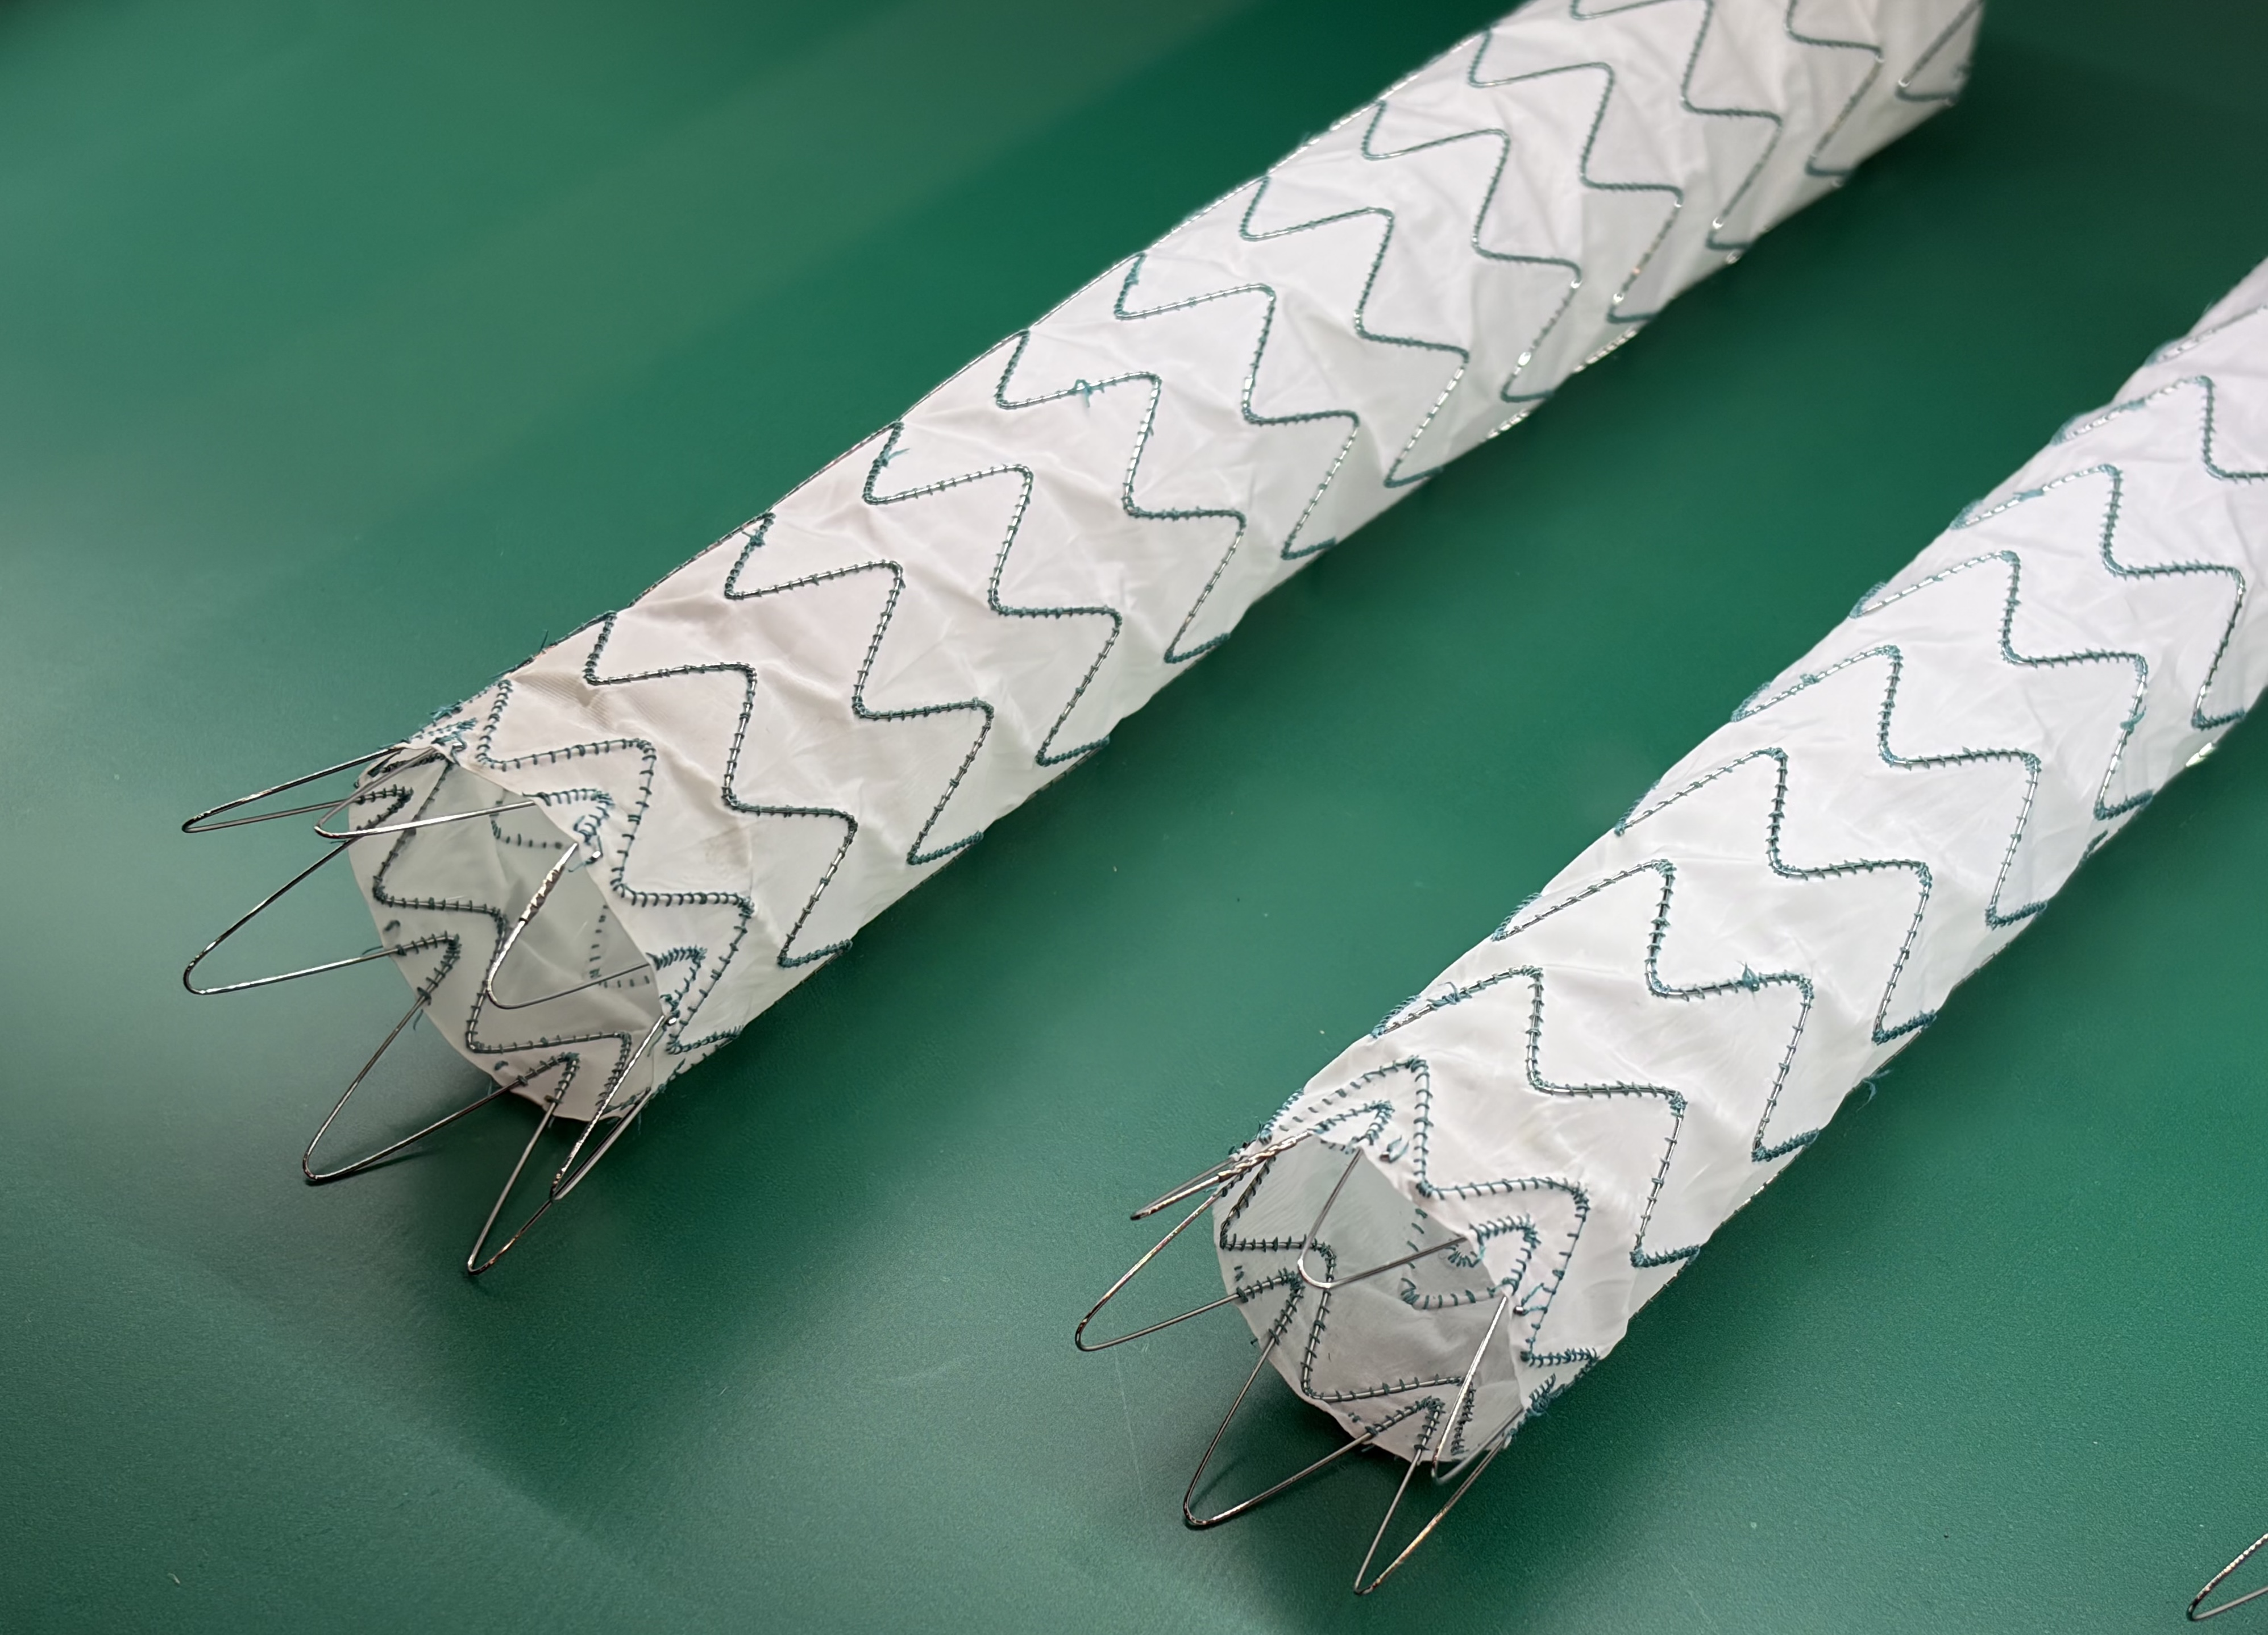
Approximately one month later, the FDA categorized this issue as a Class I recall. Even then, however, customers were not explicitly told to return the devices to the manufacturer.
Updated instructions for customers
In an updated advisory, the FDA and the manufacturer are urging customers to stop using these devices immediately and return any unused inventory to prevent them from being used by accident. In addition, customers are told to make sure any other facilities that may have received these devices are aware of these updated recommendations.
Click here to read the updated advisory.
Bolton Medical is the original manufacturer of these thoracic stent graft systems. That company is now a part of Terumo Aortic.
Michael has more than 19 years of experience as a professional writer and editor. He has written at length about cardiology, radiology, artificial intelligence and other key healthcare topics.