FDA confirms more recalls for Johnson & Johnson heart pumps due to malfunction risk
The U.S. Food and Drug Administration (FDA) has finalized three new recalls for Johnson & Johnson MedTech’s Impella RP, Impella RP with SmartAssist and Impella RP Flex with SmartAssist devices.
These heart pumps were designed to provide support to patients with acute right heart failure. According to this new warning from the FDA, some of the recordings being shared with users may not be accurate due to malfunctioning differential pressure (dP) sensors.
Unlike many recalls, no devices need to be returned to the manufacturer. However, customers are urged to read the updated the instructions for use (IFU) for these heart pumps to fully understand any potential risks.
FDA previously warned the public about this issue in February
The FDA first warned the public about this issue in February, just days after Johnson & Johnson MedTech sent an Urgent Medical Device Correction to all affected customers.
At that time, the agency was still reviewing the situation; no official recalls had been finalized. Now, however, three separate recalls have been announced. They have all been categorized as Class I recalls, which means the FDA believes using the devices without reviewing the updated IFU could result in a serious injury or death.
This safety concern has been tied to a total of 22 patient injuries. No patients have died.
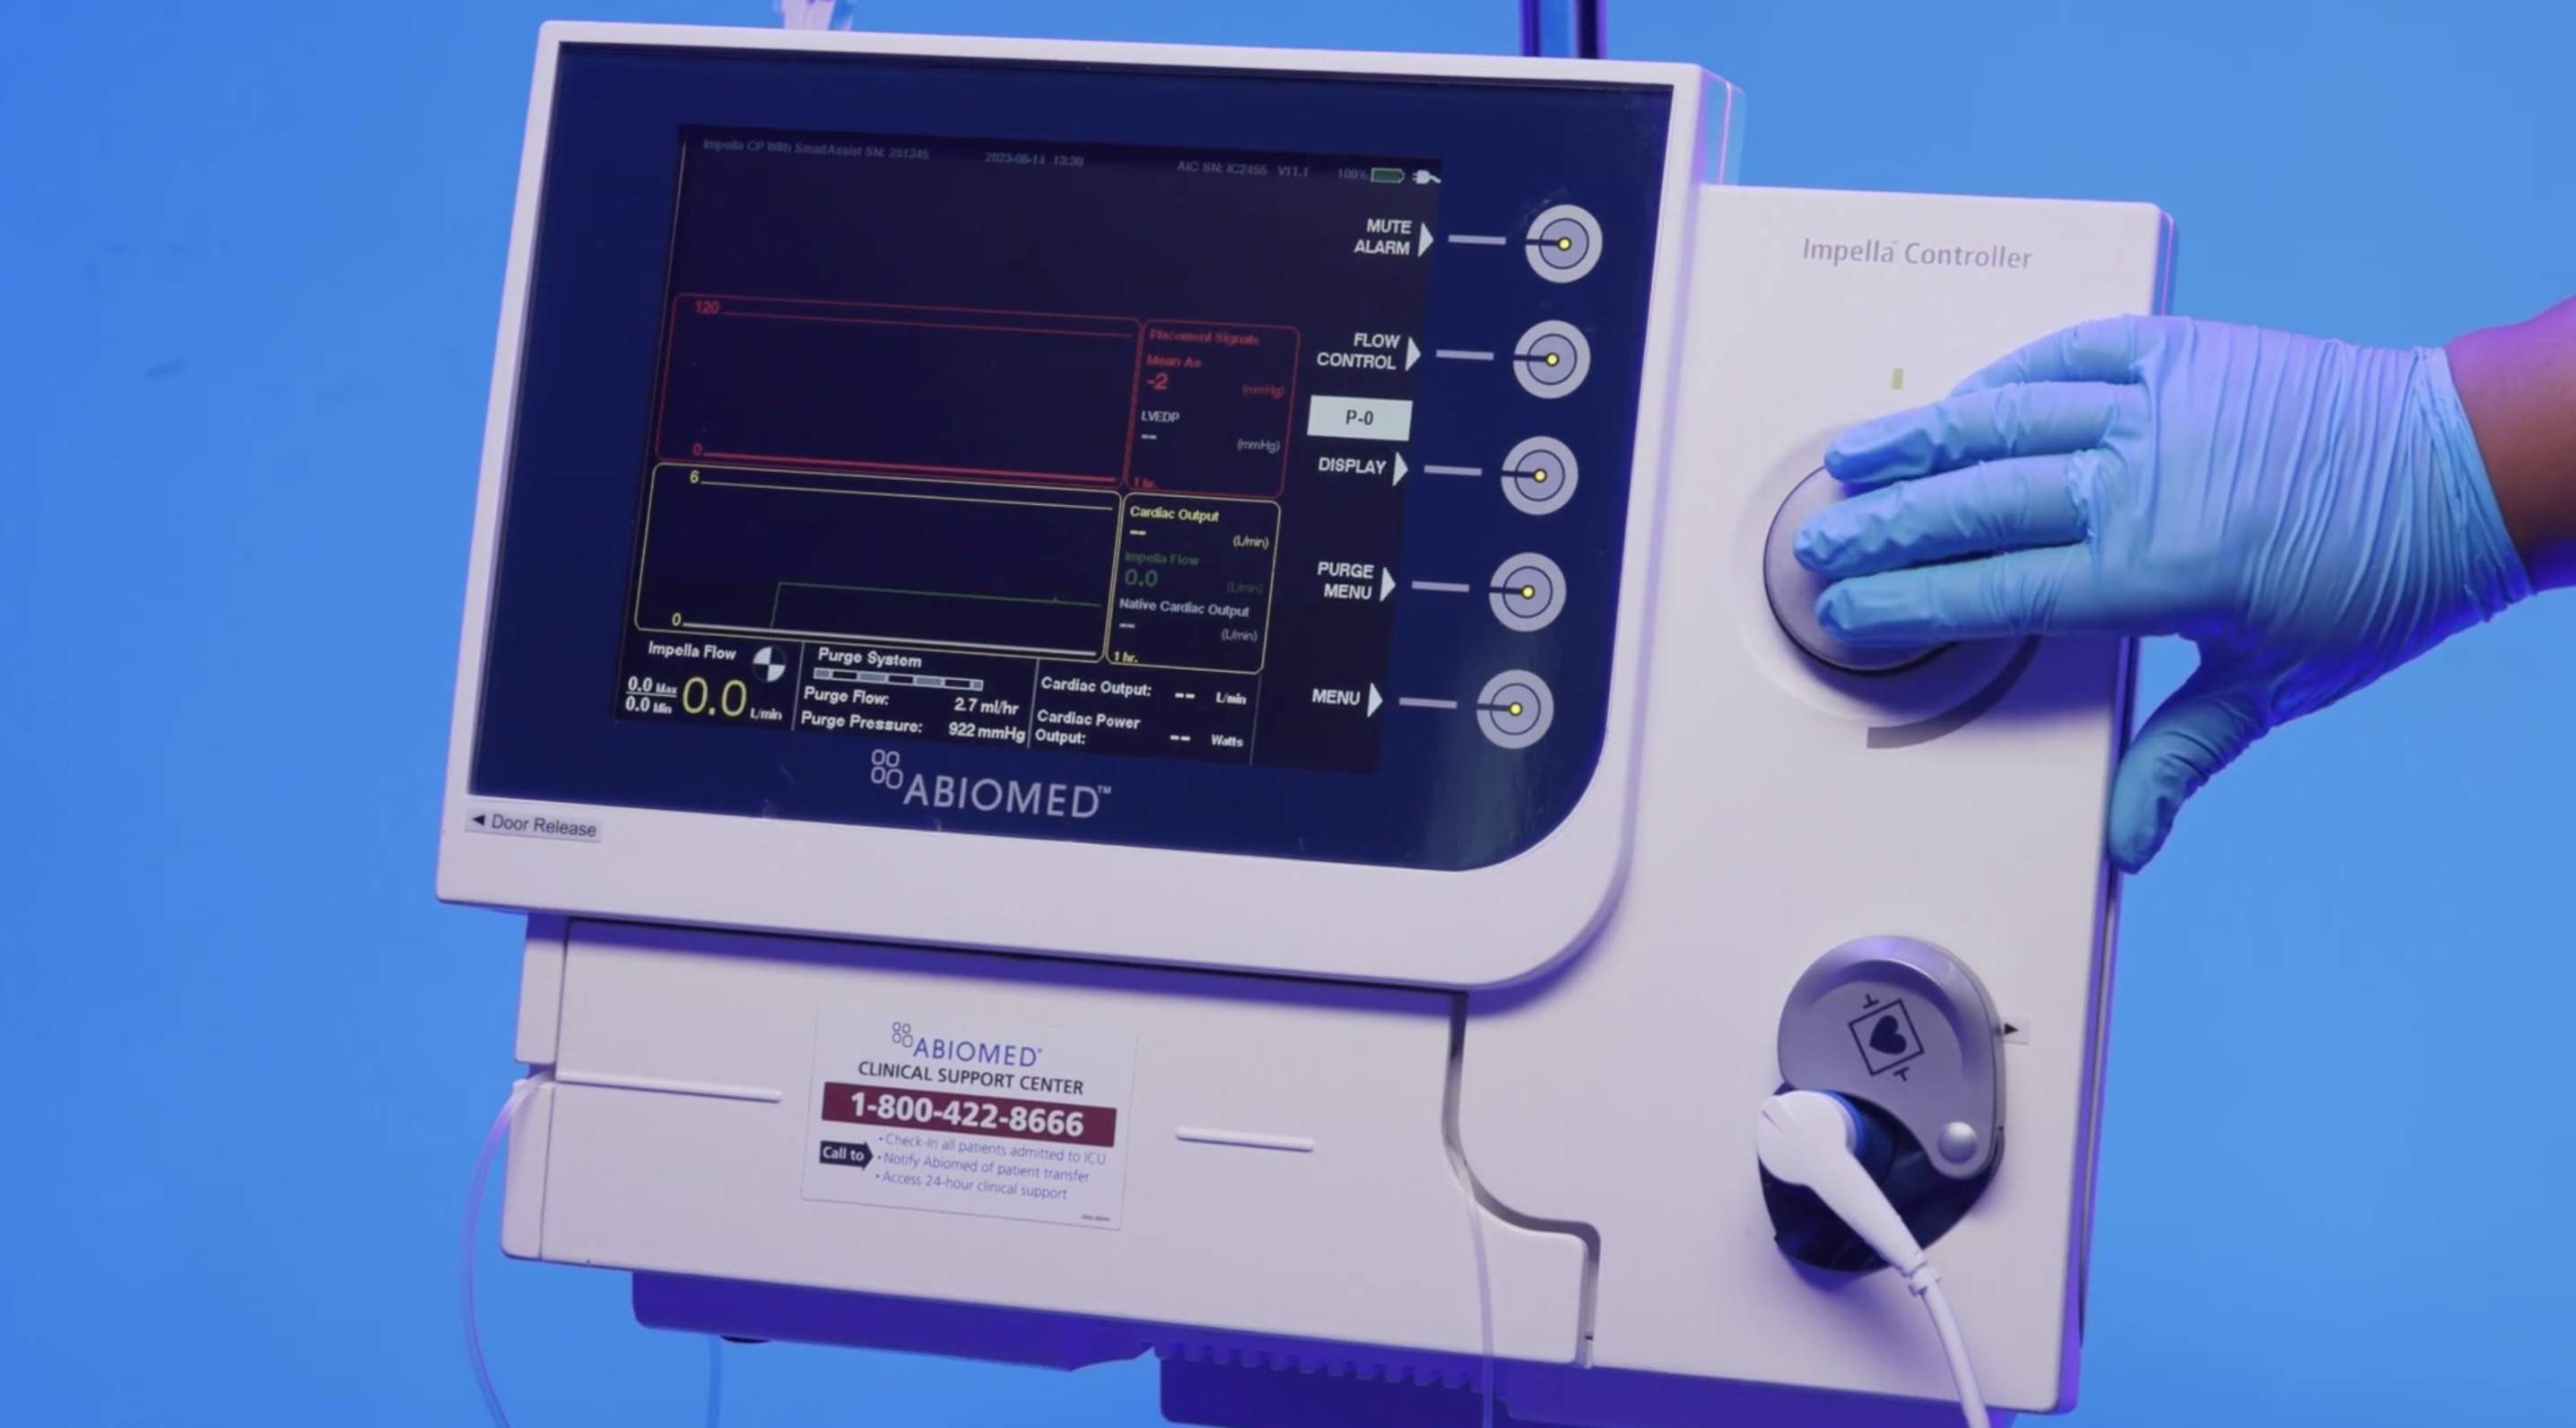
Johnson & Johnson MedTech’s Automated Impella Controller. Image courtesy of Johnson & Johnson MedTech.
Additional details about the malfunctioning differential pressure sensors
The malfunctioning dP sensors at the heart of this issue may result in values that “drift” over time, and the Automated Impella Controller (AIC) may display inaccurate information as a result. This can cause clinicians using the devices to make clinical treatment decisions they may have otherwise not made.
Johnson & Johnson MedTech has emphasized that these sensors do not impact the pump’s ability to provide patients with hemodynamic support. The company also noted that dP sensors are only present on Impella RP devices, which explains why no Impella 5.5 or Impella CP devices are included in this alert.
In the face of this issue, customers are urged to continue monitoring their patients and verifying Impella positioning prior to clinical interventions. In addition, the updated IFU document includes information that should be used to guide treatment strategies over the AIC display.
The Johnson & Johnson MedTech notification also notes that any “sudden change” in displayed flow rate should be assessed against the information in the IFU as well as temporal trends.
“An abrupt change in flow or abrupt discrepancy from the expected flow rate listed in the IFU may indicate a need to reassess pump positioning and performance, patient conditions, and clinical hemodynamics,” according to the advisory.
These recalls include 4,496 Impella RP Flex with SmartAssist devices, 291 Impella RP with SmartAssist devices and 179 Impella RP devices.
Previous safety concerns with Impella devices
Johnson & Johnson MedTech’s Impella platform has been associated with a significant number of recalls and safety alerts. In the last several months alone, AIC issues were linked to one recall in July 2025, a second recall in September 2025, a third and fourth recall in October 2025 and then a fifth recall in December 2025. In addition, the FDA just announced a potential issue with purge cassette leaks that is still under review.
Johnson & Johnson acquired Abiomed for $16.6B in 2022
The Impella line of heart pumps were previously sold under the Abiomed brand before Johnson & Johnson acquired Abiomed in 2022 for approximately $16.6 billion. In 2024, Abiomed started operating under the name Johnson & Johnson MedTech.
Michael has more than 19 years of experience as a professional writer and editor. He has written at length about cardiology, radiology, artificial intelligence and other key healthcare topics.